Selection along a gene
You have a test of selection at each codon of a gene: HyPhy's FEL or MEME, a site model from PAML, or a table of your own.
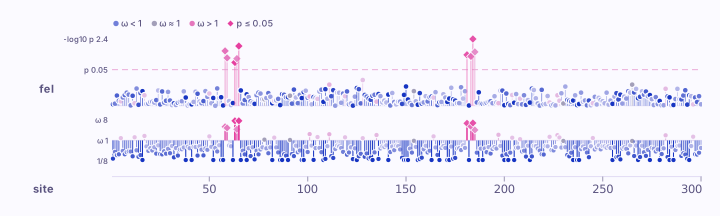
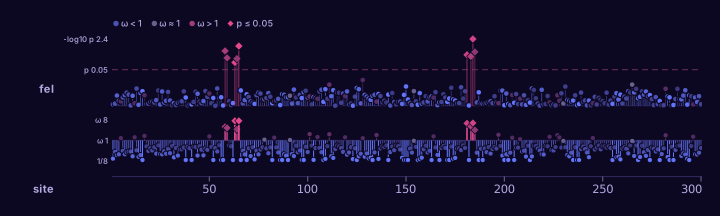
Above, the evidence at each site, as -log10 of its p-value, and the sites at
p ≤ 0.05 as diamonds. Below, its direction: ω, which is dN/dS, above 1 upwards
and below 1 downwards. The ruler counts sites from 1, as HyPhy does.
Your table
HyPhy's CSV is read as it is: alpha, beta and p-value, one row per site
in order. Any other table names its columns in a header: a site or codon
where the rows are not in order from 1, the rates as alpha and beta, dS
and dN, or their ratio as omega, and the evidence as a p-value or a
posterior. MEME's beta-, beta+ and p+ are drawn as its two rate
classes.
Change it
| To | Write |
|---|---|
| A stricter line | --threshold 0.01 |
| A table of posteriors, from FUBAR or a Bayes empirical Bayes | nothing: the header says so, and --threshold 0.95 moves the line |
| Some of the sites | site:50-200 as the place, before the file |
The example file: fel.csv. Every option:
karyon help selection, or the command line reference.