Reads, calls and genes
You have aligned reads (a BAM with its .bai), variant calls (a VCF, gzipped
or not) and an annotation (GFF3 or GTF).
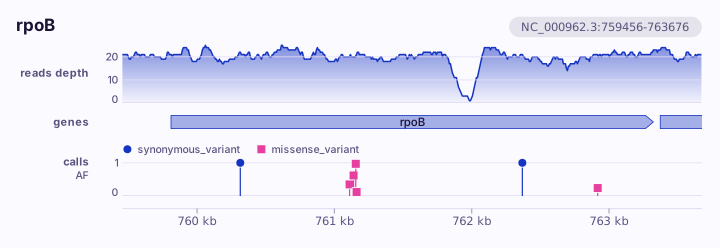
- reads depth: how many reads cover each base. The dip is a stretch no read covers.
- genes: each gene once, with its name and its direction.
- calls: each variant at its position, as tall as its allele frequency, coloured by what the annotation says it does.
Change it
| To | Write |
|---|---|
| See the reads, not their depth | --pileup reads.bam |
| See the bases, zoomed in | NC_000962.3:761,100-761,200 --pileup reads.bam ref.fa |
| Draw a wider stretch | NC_000962.3:755,000-770,000 in place of rpoB |
| Draw two genes, one under the other | rpoB rpoC in place of rpoB |
| Make a row taller | reads.bam --height 100 |
| Rename a row | calls.vcf.gz --label "variant calls" |
The example files:
reads.bam, reads.bam.bai,
genes.gff3, calls.vcf.gz and
ref.fa. Every option: karyon help coverage, or the
command line reference.